search

Chemaxon Descriptor Generation Toolkit

Chemistry calculators for increasing certainty in drug discovery

Descriptor Generation Toolkit

Assessing compounds in-silico is the most efficient way to prioritize new ideas, making sure only the best options are pursued and so minimizing effort spent on less promising compounds.

The Descriptor Generation Toolkit gives you the scientifically accurate property predictions needed to achieve this. It also allows you to generate the descriptors needed to thoroughly characterize the structures to train new models on your data.

Descriptor Generation Tools

ADMET

Chemaxon offers predictors covering key ADMET liabilities, so you can filter out weak candidates before synthesis.

  • CNS (Central Nervous System) MPO score is a composite metric combining MW, logP, logD, TPSA, HBD, and basic pKa to estimate how likely a compound is to succeed as a CNS drug.
  • Blood-brain barrier (BBB) Score is a composite metric based on aromatic ring count, heavy atom count, MWHBN (a descriptor calculated from the molecular weight and the number of H-bond donor and acceptor atoms), TPSA, and pKa that predicts whether a candidate is likely to cross the blood-brain barrier.
  • Chemaxon’s hERG predictor uses machine learning models to estimate hERG activity and classify compounds as SAFE or TOXIC, flagging risky scaffolds early in design.

Elemental Analysis

Elemental Analysis provides fundamental molecular properties for general chemistry workflows and analytical scientists.

  • Exact and nominal mass, molecular weight and atom count
  • Elemental composition
  • Molecular formulae
  • Molecular composition
  • Mass spectrum (isotope distribution) calculation

Partitioning

The partitioning of drugs in microscopic environments (e.g.: lipid bilayers of biological membranes) of different lipophilicity and hydrophilicity heavily influences its pharmacokinetic behavior. The partitioning is described by the logP and logD values of a drug, which are major descriptors in predicting ADMET properties.

The Partitioning calculator provides:

  • LogP values using scientifically sound methods
  • LogD (pH vs. logD) plot
  • HLB (Hydrophilic Lipophilic Balance) number

Solubility

Solubility in water (commonly referred to as logS) is one of the most important parameters to achieve for desired pharmacological response.

As the majority of drugs are molecules that have ionizable groups, the solubility of the compound depends highly on the pH environment. The Solubility Predictor is able to predict:

  • Intrinsic (thermodynamic) solubility in water
  • pH-dependent solubility
  • A qualitative solubility category

NMR Predictor

Nuclear Magnetic Resonance spectroscopy is the primary experimental technique to confirm and determine the chemical structure of organic compounds.

As NMR spectrometers are relatively expensive, predicting NMR spectra for a set of possible structures and comparing them with experimental data is a well-established approach to facilitate structure elucidation. The NMR Predictor is able to:

  • Predict 13C and 1H NMR spectra for molecules composed of the most frequent elements (H, C, N, O, F, Cl, Br, I, P, S, Si, Se, B, Sn, Ge, Te and As)
  • Consider H-H, H-F and C-F couplings in NMR spectrum calculation
  • Differentiate between diastereotopic protons

Structural Calculations

Structural Calculations provide:

  • Hydrogen Bond Donor/Acceptor (HBDA) count
  • 2D topological descriptors
  • 3D geometrical descriptors
  • Molecular surface calculations
  • 3D conformer generation, molecular dynamics
  • 3D Alignment
  • Different electronic property calculations

Isomers

The ability to deal with molecular structures that have identical atomic composition, but different arrangement of bonds or spatial orientation is crucial in the identification, search and property calculation of compounds. Tautomerism, stereoisomerism and resonance are examples of isomerism.

Tautomerization can affect identification, searching and physico-chemical properties of molecules. Stereoisomers have the same physico-chemical properties (e.g.: melting point, solubility), but they differ in pharmacokinetic and pharmacodynamic behavior. Resonance describes the rearrangement of delocalized electrons in the molecule, which results in a set of contributing structures of the original structure.

The Isomer calculator can:

  • Generate different tautomer sets useful for searching and tautomer handling
  • Calculate tautomer distribution and major tautomer form in water
  • Generate tetrahedral and double-bond stereoisomers starting either from a 2D or 3D structure
  • Generate resonance structures

Protonation

Ionization has an essential impact on compound behavior in nearly all protic environments. On top of its application in general and analytical chemistry, relationships to drug pharmacokinetics and pharmacodynamics are well-established. Acidic and basic dissociation constants (pKa) and the microspecies distribution at different pH values are important to quantitatively describe ionization and comprehend physical and biological processes.

The Protonation calculator includes:

  • Highly-accurate calculation of pKa values along with pH dependent distribution of relevant microspecies in water
  • Calculation of the major microspecies form at a given pH value
  • Isoelectric point calculation

Fingerprints

Molecular fingerprints are bit vectors encoding of a molecule’s structural features, used to compare, search, and cluster compounds, or to build predictive machine-learning models. Chemaxon offers the following fingerprints:

  • Extended Connectivity Fingerprint (ECFP)
  • Chemical Hashed Fingerprint (CF)
  • Pharmacophore Fingerprint (PF)
  • Reaction Fingerprint (RF)

Availability

All toolkit components are available through multiple programming interfaces, including Java, Python, REST API (JChem Microservices), .NET, and a command-line interface (cxcalc). Purchased toolkit components are also available for use in Instant JChem, JChem for Office and KNIME with the appropriate additional licenses.

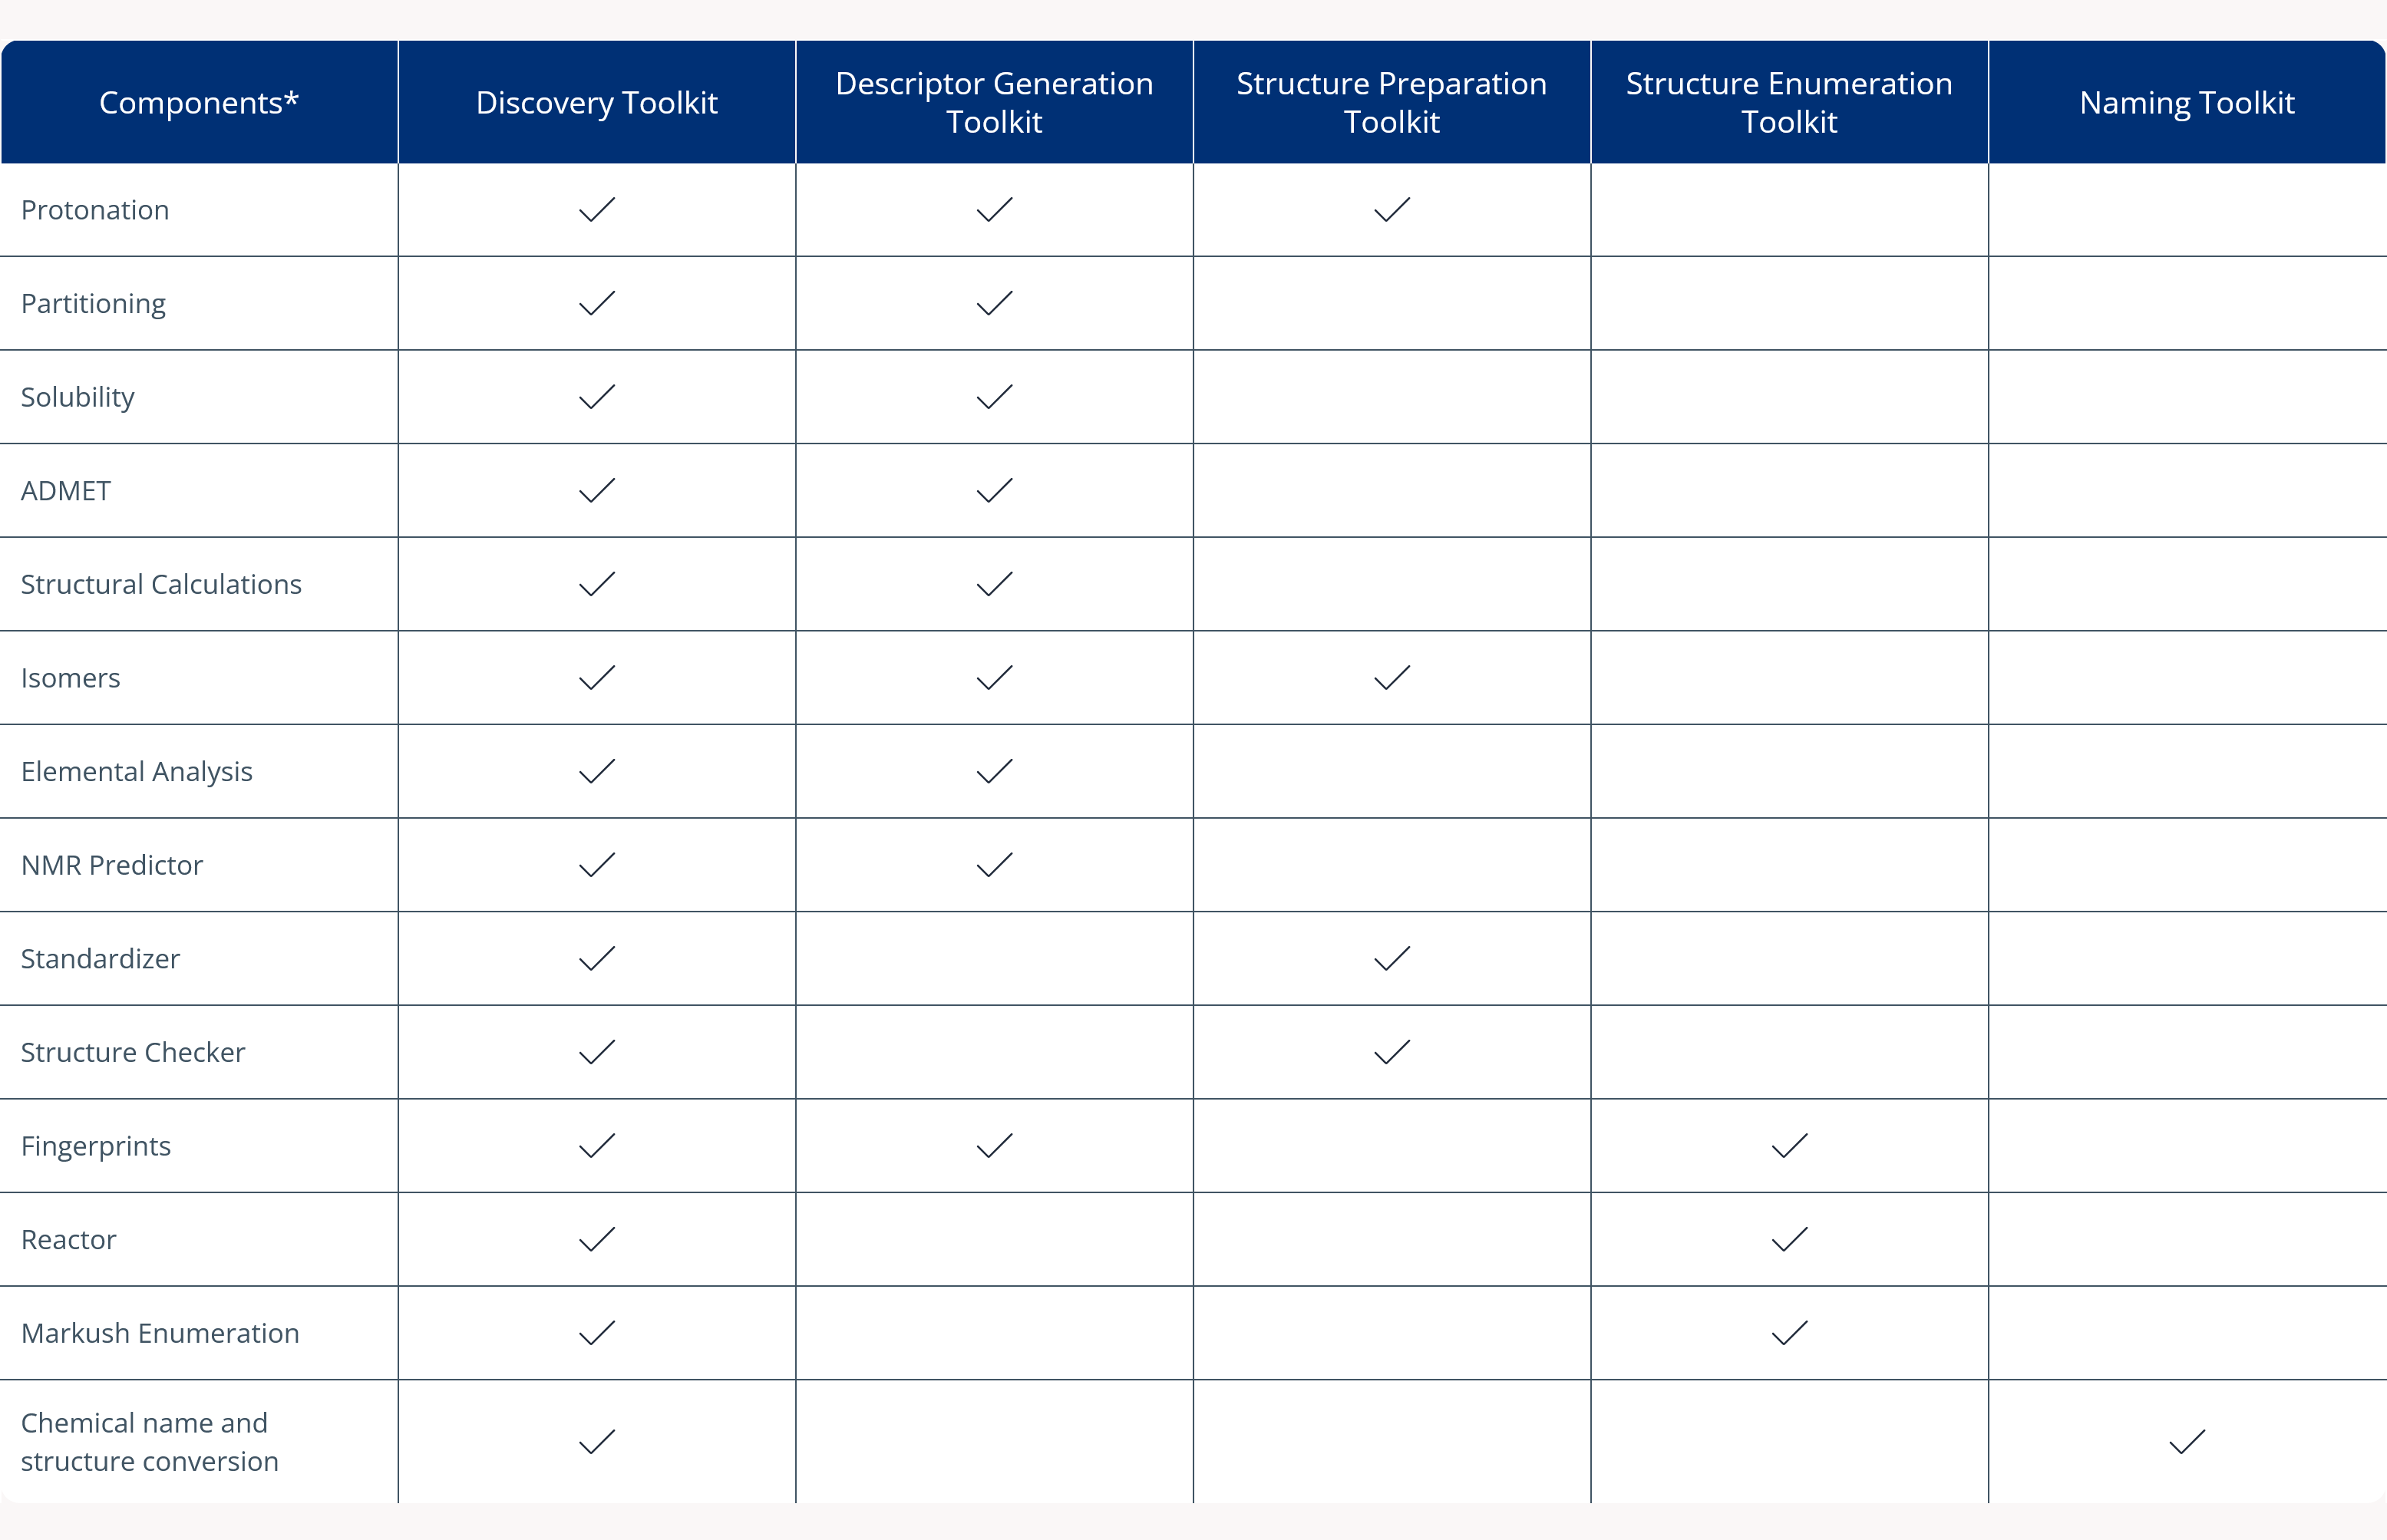
Toolkit Offerings

*For a full list of available tools and detailed descriptions, visit the corresponding bundle pages.

Workflow-Specific Toolkits

Our workflow-specific offerings are designed to serve your needs with a targeted subset of tools bundled together.

Discovery Toolkit

Read more

Structure Preparation Toolkit

Read more

Structure Enumeration Toolkit

Read more

Naming Toolkit

Read more

With Discovery Toolkit, your data is secure

Certara holds ISO 27001 certification for Certara’s Information Security Management System (ISMS). We have implemented robust security controls, undergone rigorous risk assessments, and continuously strive for improvement. Discovery Toolkit ensures full compliance with global data protection standards, offering peace of mind for sensitive analysis.

Learn more in our Trust Center

Why Certara

Certara delivers leading-edge solutions that empower scientists to make data-driven decisions, accelerating drug discovery timelines and improving outcomes.

Early drug discovery is a core area of expertise at Certara, supported by a team of highly experienced and renowned experts in the field.
The dynamic community of users from drug discovery empowers us to continuously refine our solutions to meet the ever-evolving demands of innovative research.

Using Chemaxon’s suite of tools as core components of our chemistry workflows, they save us time while supporting strong, science-driven decisions. What sets them apart is that their team is professional, approachable, and genuinely invested in helping us succeed.

Xin Zhang
Sr. Director, Cheminformatics and AI/ML
Cellarity

Book a Demo

Discovery Toolkit contains every cheminformatics toolkit we have to offer.

For more specific needs, we offer workflow-specific bundles:

Descriptor Generation Toolkit
Structure Preparation Toolkit
Structure Enumeration Toolkit
Naming Toolkit

Reach out to us

– Leave your contact information
– Summarize what you are looking for
– Our colleagues will get back to you soon